HHATL
RNA expression — cross-omicsAcross TCGA cell cohorts, HHATL RNA expression is significantly associated with the go_rna of many other GO terms, with 1,899 significant associations in total. SOFT_TISSUE shows the largest number of these associations.
The most reproducible HHATL-associated GO terms across cancer lineages are Response to chemokine, Cellular response to chemokine, and Negative regulation of mRNA metabolic process. Each is linked with HHATL in more than 4 cancer types. Because this analysis shows association rather than direction, both HHATL-to-partner and partner-to-HHATL results are reported.
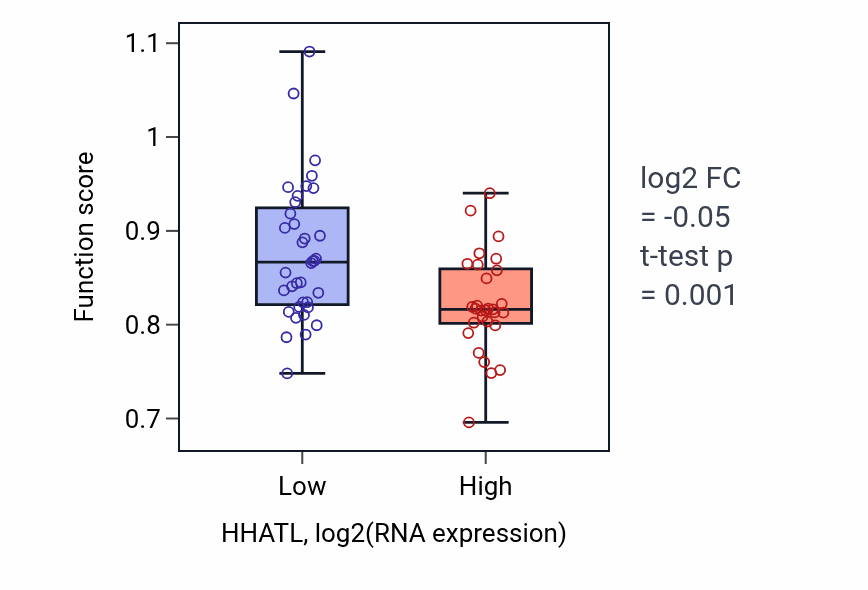
Each partner links to its own Q-omics profile. The box plot shows the strongest example, Response to chemokine grouped by HHATL-low versus HHATL-high in SOFT_TISSUE.
RNA expression associated GO terms by consensus
Ranked by combined sampling and lineage consensus. X-score (HHATL→partner) and Y-score (partner→HHATL) are standardized regression coefficients; both directions are reported because the association is undirected. p-values are from the association test.
Each partner links to its Q-omics profile. Showing the 6 strongest of 1,899 associations by consensus.